EU Authorized Representative for medical device manufacturers.
Our name goes on the label only after your file stands up to review.
Under Article 11 of the MDR or IVDR, a manufacturer with no EU place of business must appoint a single EC REP before placing a device on the market. The written mandate is effective only after the representative accepts it. MEDDEV accepts representation after reviewing the manufacturer, defined device scope, regulatory evidence, post-market controls, response readiness and liability arrangements.
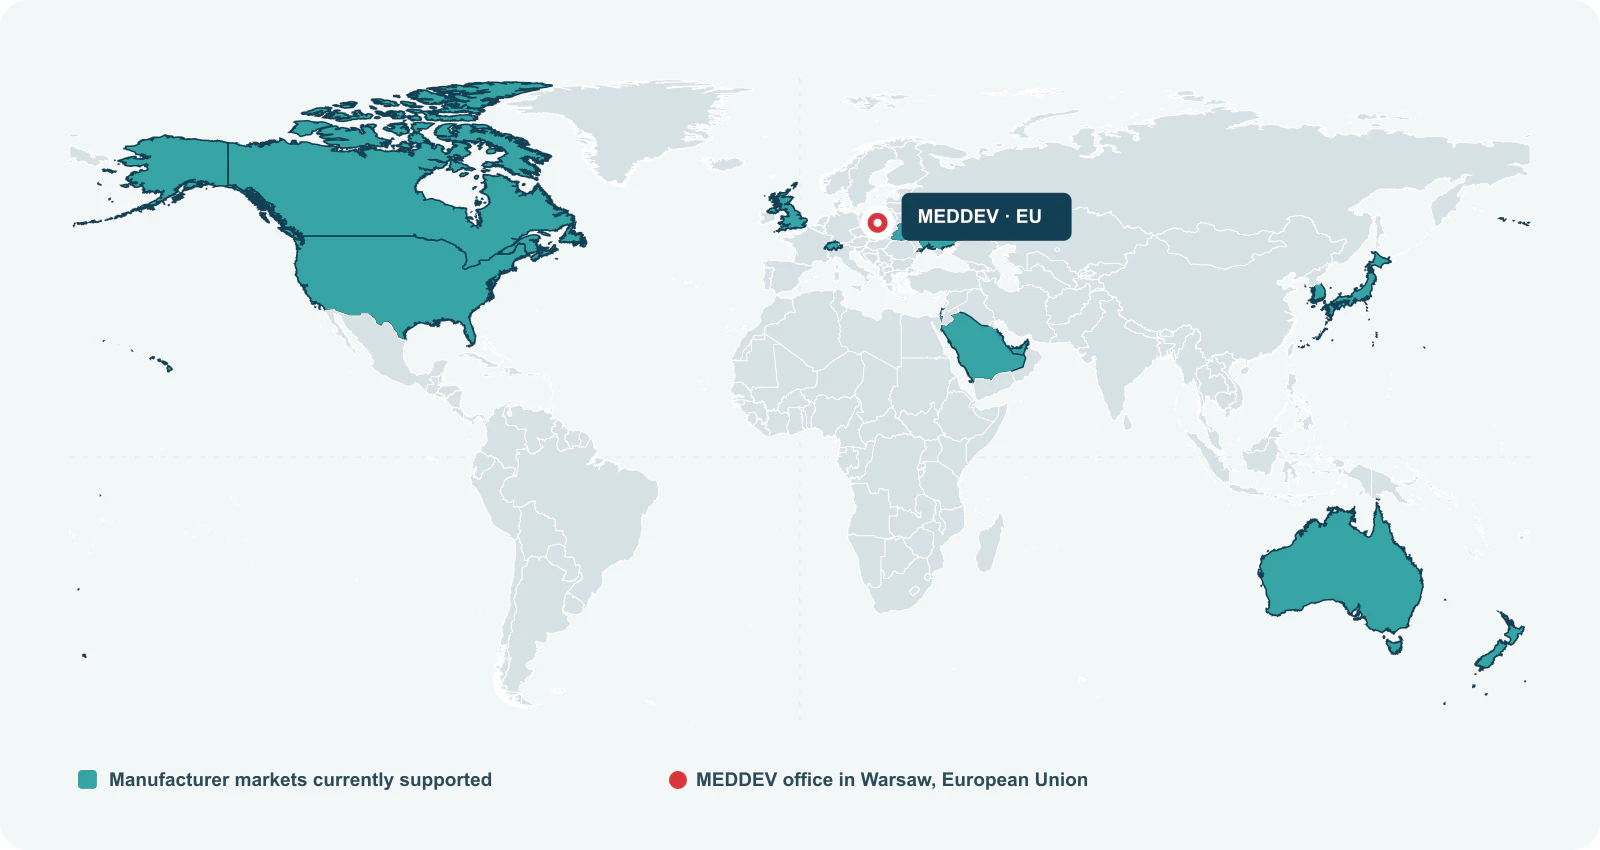
For eligible manufacturers in the UK, Switzerland, USA, Canada, Australia, New Zealand, Japan, South Korea, Saudi Arabia and the UAE — including qualifying higher-risk devices after enhanced review.
Established in the European UnionWarsaw, Poland · EC REP under MDR and IVDR Article 11
Online intakeSecure connection
MEDDEV brand serving medical-device companies since 2012MDR 2017/745 & IVDR 2017/746Mandate accepted only after review
Manufacturers who have worked with MEDDEV
MEDDEV Safety Signals · Issues 01–02
Source-backed safety reports covering Europe and the United States.
A Union route for devices designed beyond the ordinary clinic.
Trauma, evacuation, portable diagnostics, monitoring, telemedicine and resilient medical systems require evidence for the conditions in which they will actually operate. Warsaw provides an EU-established regulatory base with direct Central and Eastern European context.
Military use is not a conformity shortcut. The specialists assigned to the scope qualify the complete system, map MDR or IVDR work and assess whether a narrowly defined authority-controlled exception is legally available.
5/5 across five criteriaWDI Wise Device EuropeReadiness audit · 20155/5 across five criteriaSensiMedMedical device notification · 20155/5 across five criteriaPiórex S.A.Medical device registration · 2015
Historical MEDDEV post-project survey across regulatory services. Clients rated knowledge transfer, consultant engagement, subject-matter preparation, problem solving and overall service. The individual ratings shown here are from 2015; the summary figures cover the published set from that survey programme. Comments were optional, so most ratings carry none — and no quotation has been reconstructed or written on a client’s behalf.
Acceptance is earned, not purchased
Not every mandate is accepted.
MEDDEV accepts only defined device scopes supported by evidence, working safety controls and a manufacturer we can rely on. A commercial enquiry never guarantees appointment. Our name appears on the label only after the acceptance review is complete.
MEDDEV mandate reviewAcceptance gate
Manufacturer, device and intended purposeAcceptance criterion
Conformity route and regulatory evidenceAcceptance criterion
Safety, post-market and vigilance controlsAcceptance criterion
Communication, response and liability arrangementsAcceptance criterion
Outcome: mandate accepted, conditions issued or mandate declined.
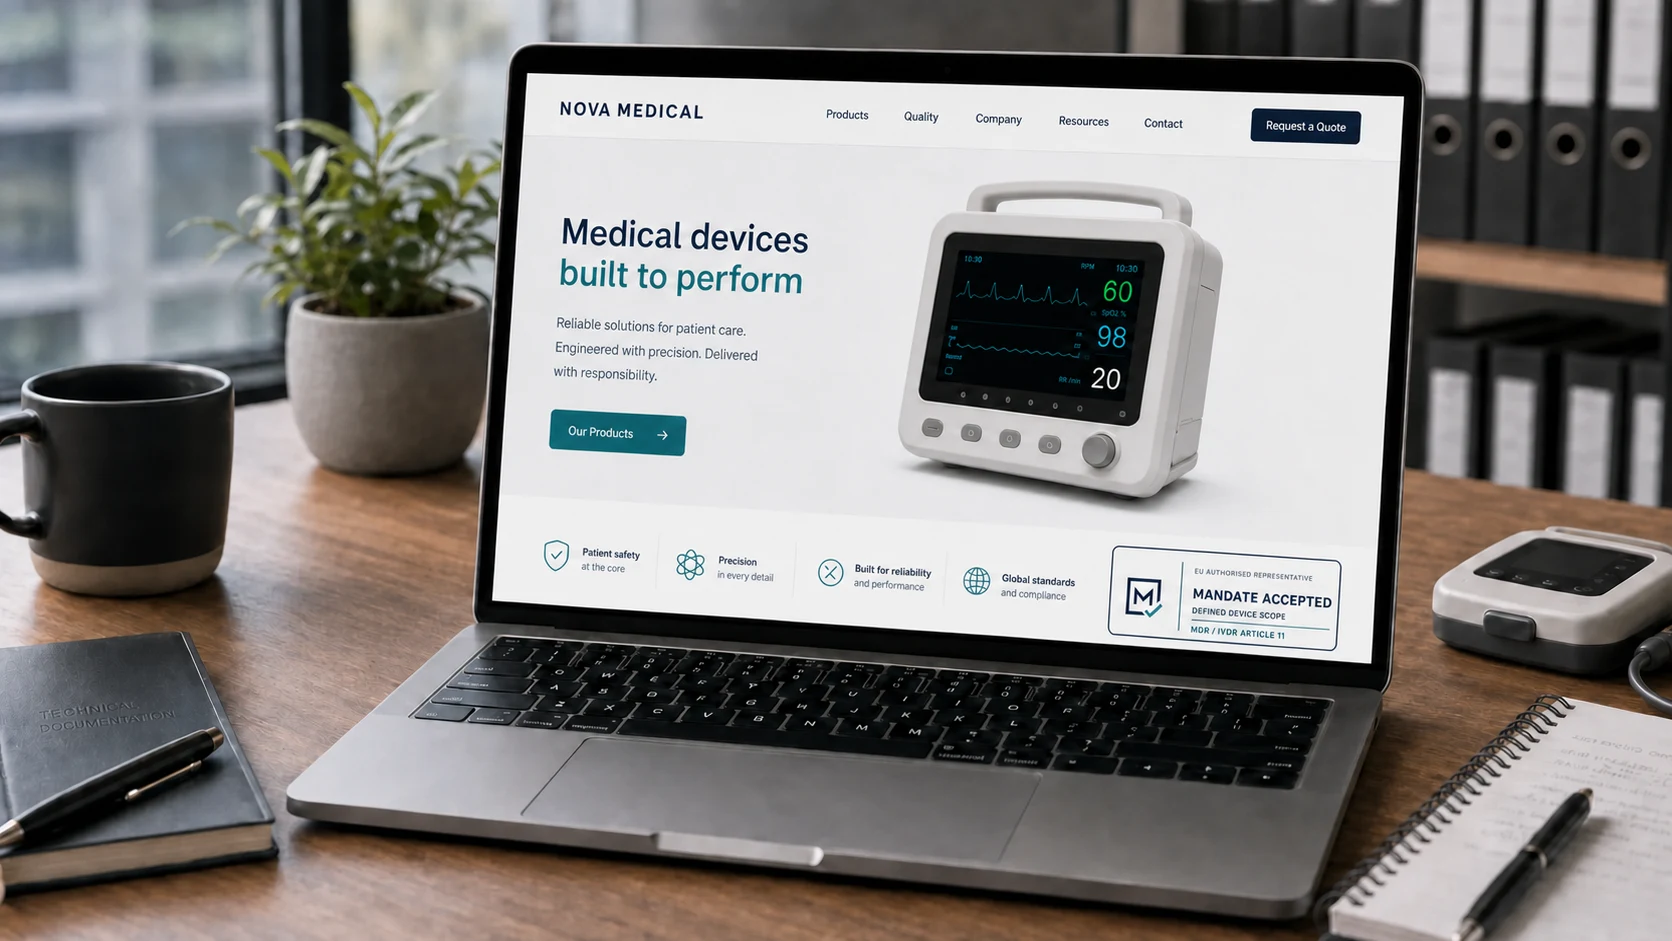
Example on a manufacturer website. The issued mark should link to the active MEDDEV mandate and its defined device scope.
A status worth displaying
Turn regulatory acceptance into a visible trust signal.
Accepted manufacturers may use the mark where professional buyers assess regulatory readiness. Each use must remain connected to the active MEDDEV mandate and its defined device scope.
Manufacturer websiteAlongside EU market-access information and the represented device portfolio.
Product catalogueOn pages covering devices included in the accepted mandate.
Distributor dossierAs a verifiable representation status during onboarding and due diligence.
Sales presentationFor procurement, partner meetings and trade-fair communication.
Use boundary: the mark must not appear as a CE mark, product certificate or safety approval. It should not be added to the regulated device label, instructions for use or sterile-packaging information panel.
Where we support manufacturers
One EU representative, wherever your company is established.
MEDDEV Compliance sp. z o.o. is established in Warsaw, Poland, inside the European Union. The mandate and ongoing work are handled remotely in English for manufacturers across the markets below.
A manufacturer established in Great Britain still needs an EU Authorised Representative to place devices on the European Union market. The EU mandate is separate from its home-market arrangements.
Japanese or South Korean market documentation can support the review, but EU evidence is assessed separately against MDR or IVDR before we accept the mandate.
Australia & New Zealand
An English-language route to EU compliance
Australian or New Zealand market documentation can support the review. It does not replace MDR or IVDR conformity, an EU representative or the applicable EUDAMED entries.
Saudi or UAE market authorization does not replace EU conformity assessment. We define the EU mandate, registration and label changes as a separate regulatory route.
Other non-EU markets
The same Article 11 rule applies
We also assess manufacturers from Ukraine, Israel and other eligible non-EU jurisdictions, subject to documentation, product-liability and sanctions screening.
Administrative appointment or active representation
A legal address is only the starting point.
Article 11 of the MDR and IVDR requires verification, record-keeping and cooperation with authorities. Because Article 11(5) can also create joint and several liability with the manufacturer when its conditions are met, the mandate needs a defined file review and response process.
When it matters
Administrative-only scope
MEDDEV active scope
When an authority requests documents
Passes the request to the manufacturer
Works from a file reviewed before the mandate and coordinates the response
During a vigilance case or FSCA
Provides the required contact point
Coordinates communication while the manufacturer investigates and decides corrective action
Technical documentation & labelling
Checks the appointment details
Reviews the agreed file scope and EC REP labelling before the mandate starts
EUDAMED registration
Leaves entries to separate providers
Coordinates actor and device entries with responsibilities defined in writing
Evidence for your EU importer
Provides appointment details
Provides verifiable registration and a clearly documented service scope
Before you enquire
Know what shapes the review and the fee.
We do not publish a one-size-fits-all price because the mandate depends on the device, portfolio and level of responsibility. The initial review identifies the scope before a fee is proposed.
Device and portfolio
Classification, IVD or medical device route, number of families, intended purpose and higher-risk features.
Evidence readiness
Technical documentation, clinical or performance evidence, quality-system records, certificates and labelling.
Ongoing responsibility
EUDAMED coordination, vigilance, authority response, importer interface, PRRC support and required review depth.
Fee principle: after the qualification review, we provide a defined scope and a proposal matched to the accepted mandate. No document upload is required in the first contact form.
01
Legal manufacturer
Confirm the entity, country of establishment, device family and intended EU market.
02
Core evidence
Have the classification rationale, technical-file index, conformity route and quality evidence available.
03
Post-market readiness
Identify complaint, vigilance, field-safety and authority-response owners before the mandate begins.
These anonymised composite scenarios reflect recurring qualification patterns. They are not attributed client case studies or performance claims; the exact scope and outcome depend on the manufacturer and device file.
01
New EU launch
Problem: home-market approval is mistaken for EU access. Work: map MDR/IVDR, labelling, EUDAMED and importer responsibilities. Typical result: a defined launch route and a documented list of open evidence.
02
Change of EC REP
Problem: the outgoing representative controls records and dates are unclear. Work: plan custody, vigilance ownership and handover steps. Typical result: a controlled transition with responsibilities recorded before the new mandate starts.
03
Higher-risk review
Problem: a Class IIb, Class III or complex IVD portfolio has unresolved evidence or liability gaps. Work: enhanced file and response-readiness review. Typical result: accept, request defined evidence, or decline before exposure is created.
EC REP mandate and supporting services
Representation plus the EU launch work around it
Start with one accountable EU Authorized Representative, then add only the supporting work your route requires: file-gap review, EUDAMED coordination, representative labelling, authority response, vigilance communication or an eligible PRRC arrangement.
A legal EU market entry
Without an EU entity of your own — we take on the mandate required by Article 11 of the MDR or IVDR.
EUDAMED registration coordinated
We coordinate actor registration, the SRN process and required device entries. The manufacturer remains responsible for the accuracy of its device data.
Label-ready compliance
Your device carries the EU representative name and address required by the applicable MDR or IVDR labelling rules — no relabelling scramble later.
A defined authority-response process
Requests for documentation or samples are assessed and coordinated without losing time on finding the responsible person.
Vigilance communication coordinated
We forward complaints and incident reports and coordinate communication with authorities. Investigation and corrective decisions remain with the manufacturer.
PRRC arranged where permitted
A separate PRRC arrangement is available for eligible micro and small manufacturers. MEDDEV also maintains the PRRC capacity required of an authorized representative.
Aesthetic surgery and energy-based devices
EU market access for a fast-moving device category.
MEDDEV supports manufacturers of surgical and aesthetic systems, including lasers, intense pulsed light, radiofrequency platforms, treatment accessories and products that may fall under MDR Annex XVI. The review starts with intended purpose, claims and classification — not with a generic registration checklist.
MDR or Annex XVI qualification and applicable common specifications
Classification, conformity route and notified-body evidence
Clinical, risk-management and post-market surveillance documentation
EC REP mandate, EUDAMED coordination and EU labelling
The obligations manufacturers outside the Union ask about most.
Each briefing states the provision it relies on and links to the official text. They are written for companies deciding whether, and in what order, to enter the European market.
EUDAMED · Article 123(3)
Device data falls due on 27 November 2026.
Four modules became mandatory on 28 May 2026. The twelve-month obligation does not cover every device, and the wording decides which.
Priority goes to complete, traceable applications with a named regulatory owner. Meeting the entry criteria qualifies a scope for a decision; it does not guarantee acceptance.
01
Verified legal manufacturer and scope
The legal entity, authorised signatory, manufacturing sites, device families, models and intended purposes are consistent and traceable.
02
Defensible EU regulatory route
The qualification, classification and conformity-assessment route are documented, with valid notified-body involvement where required.
03
Controlled regulatory evidence
The EU declaration, agreed technical-documentation index, certificates, labelling and registration data describe the same device scope.
04
Working safety and post-market controls
Risk management, surveillance, complaints, vigilance, recalls and corrective actions have current records, owners and escalation routes.
05
Responsive manufacturer relationship
Named decision-makers can provide records, samples and technical answers and can act within agreed authority-response times.
06
Liability and continuity secured
Insurance, contractual responsibility, document access, sanctions screening, change notification and mandate continuity are acceptable.
What the status means: “MEDDEV Mandate Accepted” confirms MEDDEV’s written acceptance of an EC REP mandate for the defined device scope. It is a trust signal about regulatory readiness and the manufacturer relationship. It is not a CE certificate, product approval, independent safety claim or substitute for the applicable conformity assessment.
Do I actually need an EU Authorized Representative?
If you manufacture medical devices or IVDs and have no registered place of business in the EU, Article 11 of the MDR or IVDR requires an authorized representative before the device can be placed on the EU market.
Does a Class I device still need an EC REP?
Yes, when the legal manufacturer is established outside the EU and the device is placed on the Union market. Self-certification may affect the conformity route, but it does not remove the Article 11 representative requirement.
Can FDA, Health Canada or Swissmedic approval be transferred to the EU?
No approval transfers automatically. Home-market records can support the review, but the intended purpose, classification, technical documentation, conformity route, labelling and post-market evidence must be assessed against the MDR or IVDR.
Does the Cyber Resilience Act apply to medical devices?
Medical devices and IVDs covered by the MDR or IVDR are outside the CRA scope. Cybersecurity is still a mandatory part of the MDR or IVDR compliance route, including risk management, technical documentation, updates and post-market monitoring.
Can my distributor also be my EU Authorized Representative?
A distributor and an authorized representative are separate economic-operator roles. A distributor may only act as the representative when the manufacturer gives a written mandate and the organisation can fulfil the Article 11 duties independently and continuously.
What happens if our documentation is not complete?
MEDDEV does not start a mandate on the basis of an EU address alone. We identify the missing evidence, record the acceptance conditions and either continue after the gaps are addressed or decline the mandate when the risk cannot be controlled.
Is an EC REP only a legal address?
No. Article 11 of the MDR and IVDR assigns defined verification, record-keeping, communication and cooperation duties to the authorized representative. Article 11(5) can also create joint and several liability with the manufacturer when its conditions are met. That is why every MEDDEV mandate begins with a file and liability review.
How quickly can the mandate start?
We normally confirm eligibility and the review scope on the same working day. The mandate starts after the required documentation, insurance and screening checks are complete; timing therefore depends on file readiness.
What will my EU importer be able to verify?
Before placing the device on the market, your importer must verify that the manufacturer has appointed an authorized representative and met the applicable registration obligations.
Can I switch from my current EC REP?
Yes. MDR and IVDR provide for a written changeover arrangement. We plan the transfer with the outgoing representative and manufacturer to preserve regulatory continuity.
Does the CE mark replace authorization in Saudi Arabia or the UAE?
No. Saudi Arabia and the United Arab Emirates apply their own market-access requirements. EU documentation may be used as supporting evidence where the local route allows it, but it does not replace local authorization or a locally required representative.
Can one authorized representative support both the EU and Türkiye?
For a manufacturer established outside both the EU and Türkiye, the EU–Türkiye Customs Union arrangements allow one authorized representative established either in the EU or in Türkiye for access to those markets. A manufacturer established in Türkiye does not need to designate an EU authorized representative to place devices on the EU market.
Which countries do you support?
We work in English with eligible medical device and IVD manufacturers established outside the EU, including the United Kingdom, Switzerland, United States, Canada, Australia, New Zealand, Japan, South Korea, Saudi Arabia, the United Arab Emirates, Ukraine and Israel. Every mandate starts with documentation, product-liability and sanctions screening.
Authorized or authorised representative — which spelling is correct?
Both phrases describe the same EU role. EU legislation and British English use “authorised representative”; American English commonly uses “authorized representative”. EC REP is the widely used short form.
Where is MEDDEV established?
MEDDEV Compliance sp. z o.o. is established in Warsaw, Poland, inside the European Union. We serve non-EU manufacturers remotely in English and act as their regulatory contact in the EU.
Do you work in English?
Yes — the entire mandate, correspondence and reporting are handled in English.
The people behind your mandate
Director-level governance. Named specialists for regulatory execution.
Mikołaj Junosza-Szaniawski leads mandate governance, risk oversight and international delivery. Marta Junosza-Szaniawska and Barbara Jęcka-Izdebska remain the named specialists for conformity assessment, registration and ongoing regulatory coordination.
MEDDEV Compliance sp. z o.o. · Warsaw, European Unionul. Mydlarska 47, 04-690 Warszawa, Polandbiuro@meddev.pl
Managing Director · EC REP Operations & Governance
Mikołaj brings more than 25 years of experience at KPMG, including as a Director in Audit Assurance, where he oversaw major sustainability-reporting assurance projects. He combines international project governance with a product-life-cycle perspective encompassing LCA, reuse, recyclability and environmental performance. At MEDDEV, he oversees mandate governance, risk escalation, specialist assignment and continuity of manufacturer communication.
25+ years at KPMGDirector, Audit AssuranceMBA · Carlson School of ManagementSpecialist LCA training · MITESG lecturer · Kozminski UniversityProgramme manager · EPD Polska
One accountable contact, specialist depth when the device requires it.
The manufacturer does not need to coordinate a loose group of advisers. The named coordinators define the review scope, assign the relevant specialists and remain accountable for communication and the agreed regulatory work.
Regulatory affairsQuality systemsClinical evaluationPhysiciansBiomedical engineeringSoftware and cybersecurityAesthetic devicesPost-market surveillance
Marta Junosza-Szaniawska
Conformity assessment & EC REP mandate review
Marta coordinates the initial regulatory assessment, defines the file-review scope and confirms whether MEDDEV can accept the representative mandate.
With your permission, we use Google Analytics, Google Ads conversion measurement and Microsoft Clarity to understand how the website is used. Form contents are excluded from recordings. Privacy policy.

 Online intake Secure connection